Icelandic Medicines Agency
Safety information
The marketing authorisation of certain medicines is subject to conditions concerning safe use. These are specific measures that the marketing authorisation holder must implement in order to minimise risk and/or ensure proper efficacy when the medicines are used.
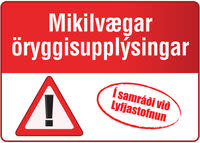
In many cases, these include the publication of safety information for healthcare professionals and patients. All material published in connection with such conditions is reviewed by the Icelandic Medicines Agency (IMA). Safety information published in connection with such conditions is identified with a special logo for important safety information, see below.
It is essential that healthcare professionals who prescribe or administer the medicine concerned, and patients/relatives who use the medicines, familiarise themselves with the relevant instructions relating to the published safety information.
IMA allows two weeks to verify that all relevant documentation accompanies an application for the assessment of safety information for medicines.
IMA can then take up to 60 days to complete the review of the safety information.
1. General requirements
Safety information is part of the risk management plan (RMP) under the conditions of the marketing authorisation and is classified as additional risk minimisation measures (aRMM). Safety information shall comply with the requirements set out in Annex II D and/or the RMP, as well as GVP V Rev 3, GVP XVI – Risk minimisation measures and GVP XVI Addendum II – Methods for evaluating effectiveness of risk minimisation measures.
The Icelandic Medicines Agency (IMA) reviews the safety information against the requirements set out and the circumstances in Iceland, and proposes changes as necessary. When new conditions concerning safety information are imposed after the marketing authorisation has been granted, or if conditions are changed, the marketing authorisation holder (MAH) is responsible for fulfilling the new/amended conditions.
1. 1 Generic medicines
In the case of a generic medicine where safety information already exists for the reference medicine (the originator or another generic medicine), the wording, objectives, content and presentation of the safety information for the generic medicine shall not differ from the safety information for the reference medicine unless there is clear justification for doing so. This is to minimise the burden on the healthcare system, ensure consistency and increase patient safety.
Marketing authorisation holders are encouraged to prepare joint safety information for the same active substance when the requirements for the safety information are comparable. When joint safety information is prepared for more than one proprietary medicinal product, only the active substance of the medicine may be referred to, and the product names may not be used.
1.2 User testing
IMA may require that safety information be user-tested with healthcare professionals and/or patients. The purpose is, among other things, to determine whether recipients understand the safety information and whether it is user-friendly.
Marketing authorisation holders are encouraged to user-test the safety information whenever possible.
It should be noted that it can be helpful to consult an expert in the relevant field when preparing the safety information.
1.3 Distribution plan
The marketing authorisation holder/local representative must submit a distribution plan, which IMA assesses. The distribution plan is part of the application form. For each component of the safety information, the following must be specified (note that the cover letter is one of the components and must therefore be specified in the plan):
Recipients: E.g. whether the component is intended for patients, doctors (by specialty), pharmacists, etc.
Distribution channel: It must be specified how the component will reach the recipients.
Number of copies to be distributed: Applies primarily to distribution on paper. It must be specified how many copies will be distributed, and this number must be based on a realistic estimate of the use of the medicine and the purpose of the component.
A minimum of three copies of patient cards shall be distributed per recipient. If considered necessary, however, more should be distributed, as appropriate.
Distribution of material: The planned date of distribution of the safety information must be specified here.
Redistribution of material: It must be specified here whether the educational material is planned to be redistributed regularly and, if so, how, or whether the material will be redistributed as needed and, if so, how this will be done. In the case of redistribution, no changes may be made to the component.
Information on user testing: It shall be specified here whether any user testing has been carried out on the component.
If the safety information is updated, an updated distribution plan must be sent to IMA.
The applicant shall complete the distribution plan that is part of the application form for the assessment of safety information, or submit a distribution plan in a separate document that meets all the same requirements. An application is not considered valid unless a completed plan is enclosed.
1.4 Distribution plan for patient cards in packs
It is not necessary to send IMA an application for the review of safety information that consists solely of a patient card that is part of the medicine pack and the approved product information. Nevertheless, a completed distribution plan for safety information must be sent to IMA when changes are made to patient cards that are part of a medicine pack, or when a patient card is added to the pack of a medicine already on the market.
The distribution plan must include information on when packs with the updated patient card are expected to reach patients and a plan for how the patient card will be distributed until then. The plan must be realistic in view of the stocks of older packs held by wholesalers/pharmacies and the sales figures of the medicine. This is to ensure that patients receive the required safety information.
1.5 Data protection
Laws and rules on the processing of personally identifiable information must be taken into account; see further on the website of the Icelandic Data Protection Authority (Persónuvernd). However, IMA does not review information concerning data protection presented in the safety information. Such information is the responsibility of the MAH/local representative.
1.6 IMA's review and distribution of safety information
New safety information shall always be sent to IMA for approval. However, this does not apply when the safety information consists solely of a patient card that is part of the medicine pack and the text of the patient card is part of the product information of the medicine.
When a substantive update is made to safety information, the safety information shall be sent to IMA for approval. The application form shall explain what changes are being made to the safety information, and the changes shall be highlighted in the documents being updated. An application for the review of safety information shall not be sent to IMA when the safety information is to be redistributed. Redistribution is permitted in accordance with the approved distribution plan, but no changes to the components are permitted, including the date in the cover letter. IMA does not accept applications for the review of educational material if no substantive changes have been made.
IMA reviews safety information for medicines that are to be placed on the market in Iceland. In the case of pharmaceutical forms, where an update to the safety information concerns new pharmaceutical forms that will not be marketed in Iceland, the safety information shall not be updated nor shall IMA's review be requested, unless there is clear justification.
Distribution of safety information
Material intended for patients – The aim of distributing safety information is to ensure that all recipients have access to the relevant components, and that the safety information reaches recipients in the way that is most suitable and best ensures that the purpose of the safety information is achieved.
IMA always requires that components intended for patients and/or carers be distributed on paper.
Material intended for healthcare professionals – IMA supports electronic distribution by email of safety information intended for healthcare professionals, with simultaneous publication of the safety information in the Medicinal Product Information Catalogue (Sérlyfjaskrá).
In the case of checklists or forms to be completed, it is generally expected that the material will be distributed on paper or that the forms can be completed electronically.
In the case of generic medicines where the safety information is fully consistent with the safety information for a reference medicine that has already been distributed and is published in Sérlyfjaskrá, publication of the safety information in Sérlyfjaskrá may be considered adequate distribution. However, this does not apply to material intended for patients.
If the applicant wishes to publish safety information on its own website, this must be specified in the application form. The website may then contain only the safety information. The applicant may not refer to additional information or websites that are not part of the safety information.
2. Application for the assessment of safety information
IMA allows 60 days to review safety information from the date of receipt by IMA. The following guidelines shall be followed when an application for the assessment of safety information is sent to IMA.
The applicant shall send IMA a completed application for the assessment of safety information. To ensure that a valid version of the form is used, it shall be downloaded from IMA's website each time an application is sent.
Documents that IMA needs to assess shall be in WORD format (.doc/docx).
The approved current conditions concerning additional measures to minimise the risks associated with the use of the medicine (Annex II and/or RMP) must be enclosed.
IMA encourages the use of the name of the active substance in the safety information instead of the proprietary name wherever possible. This is intended to cause as little confusion as possible for patients using generic medicines.
Only safety information in Icelandic shall be sent, not a foreign-language version or reference. IMA may request the foreign-language reference if necessary.
Safety information shall not contain or be linked to advertising, either directly or indirectly.
If images (photographs, drawings, graphs) are planned in the safety information, they must be included in the material when it is sent for review. Images may not be added after IMA's review.
Medicine logos are generally not permitted unless the applicant provides valid justification. The logo of the marketing authorisation holder or local representative may be used in one place on each component, on the front or back page, but care shall be taken that it is not larger than the title of the safety information.
Use of the registered trademark is only permitted the first time the name of the medicine appears.
IMA inserts a date in the form "Samþykkt af Lyfjastofnun í <mánuður><ár>" (Approved by IMA in <month><year>), which confirms that the safety information meets the conditions of the marketing authorisation. This sentence shall, as a minimum, appear on the first page of each component. The marketing authorisation holder may not insert or change the date but may choose where the date is placed in the components.
3. Guidelines on the content, layout, distribution and publication of safety information
3.1 Cover letter
A short cover letter to the recipients of the safety information must always be enclosed and shall contain at least the following:
The logo identifying important safety information.
A concise description of what the safety information is about, i.e. which risk the safety information is intended to address. The legal requirements for safety information shall not be explained.
The black triangle ▼, where applicable, together with the accompanying information. The black triangle shall appear next to the name of the medicine/active substance the first time it is used.
The requirement to report all adverse reactions to IMA. The following text may be used: Heilbrigðisstarfsmenn eru minntir á skyldur sínar til að tilkynna allar aukaverkanir sem grunur er um að tengist lyfinu. Hægt er að tilkynna til Lyfjastofnunar: Lyfjastofnun eða í gegnum eyðublað (Tilkynning um aukaverkun) sem er að finna í Sögu. (Healthcare professionals are reminded of their obligation to report all suspected adverse reactions associated with the medicine. Reports can be made to IMA: Lyfjastofnun or via the form (Tilkynning um aukaverkun) available in Saga.)
All components shall be listed in the cover letter (ensure that titles are consistent), together with where the components can be accessed.
If the components have been updated, the main changes made shall be specified.
If a previous version of the safety information has been distributed to recipients (in print or electronically), the cover letter shall remind recipients to dispose of the older version of the material.
The cover letter shall specify the recipients in accordance with the distribution plan for the material, together with a recommendation to inform other healthcare professionals of the safety information. It is the applicant's responsibility to propose recipients, which IMA assesses.
The following sentence or a similar one may be used (depending on which recipients are relevant):
Viðtakendur þessara öryggisupplýsinga eru: Allir sérfræðingar í xxx (sérgrein) og lyfjafræðingar í lyfjabúðum/sjúkrahúsapótekum. Viðtakendur eru hvattir til að láta aðra heilbrigðisstarfsmenn vita um innihald upplýsinganna eftir því sem við á. (The recipients of this safety information are: All specialists in xxx (specialty) and pharmacists in community pharmacies/hospital pharmacies. Recipients are encouraged to inform other healthcare professionals of the content of the information as appropriate.)
3.2 Components of safety information
All components of safety information shall, as a minimum, contain the following:
The logo identifying important safety information shall appear on all components.
a. IMA encourages placing the logo on all patient cards wherever possible, including patient cards that are part of the product information of the medicine.
b. If there is no space at all for the logo on a patient card sent to IMA for review, the applicant may request an exemption.
If the black triangle ▼ applies, it must appear on the material together with the accompanying information. The black triangle shall appear next to the name of the medicine/active substance the first time it is used.
The reader shall be referred to the SmPC/package leaflet, as appropriate, for further information.
The wording shall be consistent with the approved product information of the medicine, the Summary of Product Characteristics (SmPC) and the package leaflet.
A version number/identifier shall appear on all pages of the safety information.
3.3 Special additional guidelines for patient cards:
The key objective of patient cards is that patients carry them with them. They must therefore be of a suitable size (e.g. fit in a wallet) and printed on paper suitable for such use. If the patient card is included in the medicine pack, the patient must be able to identify it easily (patient cards shall not be printed on standard package leaflet paper or printed as part of the package leaflet).
IMA strongly requests that all patient cards be published on www.serlyfjaskra.is, see guidelines on the publication of safety material.
4. Reviewed safety information
When IMA has completed its review, it sends the safety information with proposed changes/corrections and comments to the applicant.
If IMA states that no further review is needed, the safety information shall not be sent back to IMA.
When this is stated, IMA has assessed the changes required as minor, and it is then the applicant's responsibility to address the comments. IMA will not review the safety information again in such cases.If IMA requests that the updated material be resent, this will be stated, and a response shall be provided within 30 days. In such cases, IMA reviews the responses and changes submitted.
If the applicant disagrees with IMA's comments/changes, IMA shall be informed within 30 days.
IMA adds a V-number associated with the medicinal project to the cover letter. The applicant may not change the V-number.
5. Distribution of safety information
The logo identifying mailings with important information shall be placed on envelopes when the safety information is sent by post, and in a prominent place near the top of the email if the information is sent by email.
PDF documents for publication/distribution must be searchable.
When safety information is distributed, no other material may accompany it.
Safety information intended for patients must always be sent in print to the relevant recipients, even if it is also available on www.serlyfjaskra.is.
An exception to this is when patient cards are included in the medicine pack; in that case, however, healthcare professionals must be informed where the card can be found and what needs to be reviewed with the patient (e.g. recording information on the card). IMA requests that patient cards included in the medicine pack be published in Sérlyfjaskrá.
6. Publication of safety information in Sérlyfjaskrá - the Medicinal Product Information Catalogue
IMA encourages the publication of safety information in Sérlyfjaskrá. See guidelines below.
7. Publication of safety information on the marketing authorisation holder's website
If safety information is to be published on the marketing authorisation holder's website, the following shall be observed:
a) Distribution of safety information via the website shall be approved by IMA.
b) A link to the page shall be included in the application form for the assessment of safety information.
c) The website may not contain links or references to other documents or websites unless IMA has approved this.
d) The website may not contain references to or information about medicines that are not on the market in Iceland.
1. General
Please note that only safety information approved by the Icelandic Medicines Agency (IMA) is published on www.serlyfjaskra.is.
Safety information for new medicines: Published at the same time as the medicine is published in the Medicinal Product Information Catalogue (Sérlyfjaskrá). IMA points out that safety information is displayed in alphabetical order in Sérlyfjaskrá. IMA uses numerical ordering to ensure that documents are displayed in the order requested.
Updated safety information: As a rule, published in Sérlyfjaskrá every working Monday, and a request for publication must therefore be sent to IMA at least 7 days before the requested publication date.
Once the safety information has been published, the applicant is responsible for checking on www.serlyfjaskra.is that everything is as it should be. If anything has gone wrong with the publication, the applicant contacts IMA.
The following are general rules to bear in mind:
2. File names
The file name must describe the content of the file (note that this applies only to the name of the file, not the title that will be published, and is intended to facilitate the filing of documents at IMA).
The file name may not exceed 50 characters.
No symbols may be used in the name (except underscore _).
No spaces may be used in the name.
No Icelandic-specific characters may be used in the name.
Example of a file name: Fraedslubaeklingur_fyrir_heilbrigdisstarfsmenn (Educational brochure for healthcare professionals).
3. Title or name of the component as published
The title or name of a component displayed on www.serlyfjaskra.is (which is clicked to open the material) may be up to 50 characters. The published title or name of the component does not need to be the same as the file name. If it proves difficult to convey the title properly in 50 characters, a field is available for adding explanations below the title (comments field).
Titles of safety information are displayed in alphabetical order unless numbers are used at the beginning of the titles for publication. IMA uses numerical ordering to ensure that educational material is displayed in the order requested by the applicant.
4. Email/request
The following table shall be used when sending a request for publication in Sérlyfjaskrá:
Month of publication: |
|---|
V-number: |
Order | Title as published in Sérlyfjaskrá | File name | Version number | Replaces (if applicable) |
|---|
Requests for publication shall be sent by email to IMA's email address: lyfjastofnun@lyfjastofnun.is.
The text in the subject line shall be „Öryggisupplýsingar fyrir [nafn lyfs] til birtingar þann [dagsetning birtingar]“ (Safety information for [name of medicine] for publication on [date of publication]).
All documents for publication must be attached to the email.
The email must state (see table above):
The V-number of the application.
The requested date of publication (in accordance with section 1 above)
All medicines concerned (name of medicine, strength and form)
Which version is concerned (example: New material, version 1, or updated material, version 3).
If updated material is to replace material already published, it must be specified exactly which components are concerned, which material is to be removed from publication and which material will replace it.
Numerical ordering is used to ensure that documents are displayed in the order requested. If a number is not used at the beginning of the title for publication, the documents are displayed in alphabetical order.
5. Video/audio files
IMA can publish safety information as a video or audio file on www.serlyfjaskra.is. The file shall be submitted by email; if the file is large, reference may be made to a website where it can be found.
The applicant must ensure that video and audio files do not exceed 100 MB.
6. Application for the withdrawal of educational material from the Medicinal Product Information
The applicant shall send IMA a request for the withdrawal of safety information from the Medicinal Product Information by email to lyfjastofnun@lyfjastofnun.is.
The text in the subject line shall be „Umsókn um niðurfellingu öryggisupplýsinga fyrir (nafn lyfs)“ (Application for the withdrawal of safety information for (name of medicine))
The email must state
All medicines concerned (name of medicine, strength and form)
Which component is to be removed from publication and when (the exact same title as published shall be used)
Updated risk management conditions must accompany the request in support of it (RMP/Annex II)
The guidelines on the preparation of safety information and the guidelines on the publication and withdrawal of safety information were last updated in December 2024.
Applying for the assessment of safety information
Application form for the assessment of safety information (docx)
Marketing authorisation holders are specifically advised to always use the current application form for the assessment of safety information. The form currently in force was updated in December 2024. Please do not use older versions of the same form when applying for the assessment of safety information.
Labelling of safety information for healthcare professionals
The Icelandic Medicines Agency (IMA) and Frumtök (the Icelandic Association of the Pharmaceutical Industry) have jointly had a logo designed to be printed or affixed on mailings to doctors and other healthcare professionals when the material for distribution is required by, or specifically approved by, IMA.
On the one hand, this concerns safety information that marketing authorisation holders are required to prepare under the conditions of a medicine's marketing authorisation and/or commitments in the risk management plan (RMP), and which concerns important aspects of medicine safety. On the other hand, it concerns Direct Healthcare Professional Communications (DHPC).
IMA and Frumtök have concluded an agreement on the use of the logo, which marketing authorisation holders in Iceland are encouraged to join; by doing so, they undertake to always use the logo to identify such mailings.
IMA recommends that the logo also be used in letters to healthcare professionals that are reviewed by IMA. This applies both to individual letters (DHPC) and to cover letters accompanying specific safety information. Nevertheless, the logo shall always be used on envelopes, whether or not the letters themselves are identified in the same way. Please note that the logo may only be used for material distributed as required by, or with the specific approval of, IMA.
Participation in the agreement is open to all marketing authorisation holders in Iceland
All marketing authorisation holders in Iceland are welcome to join this agreement. Requests to join must be sent to IMA at lyfjastofnun@lyfjastofnun.is
Abbott
Abbvie
Accord
Actavis
Actelion Registration
actrevo GmbH
Advanced Accelerator Applications
Alcon
Alfasigma
ALK-Abelló
Allergan Norden
Allergan Pharmaceuticals International Ltd.
Allergen Pharmaceuticals Ireland
Alvetra GmbH
Alvogen ehf.
Amgen
Angelini Pharma S.p.A.
Arthropharm Europe Limited
Astellas
AstraZeneca
Baxter
Bayer
Bayer Animal Health GmbH
Bene-Artzneimittel GmbH
BGP Products AB
BGP Products Aps
BGP Products Ltd
Biogen Idec
Bioglan Pharma AB
BioMarin International Limited
BMS
Boeringer Ingelheim
CCS
Celgene
Celltrion
CEPHLAPHARM Registration GmbH
Ceva Sante Animale
Chanelle UK Limited
Cheplapharm Arzneimittel
Chiesi Farmaceutici SpA
Clintec Parenteral SA
Colgate Palmolive A/S
CSL Behring AB
Daiichi Sankyo Europe GmbH
Dentsply De Trey
Desitin Arzneimittel
Diurnal
dne pharma as
Dopharma Research B.V.
Eisai
Elan Pharma International Ltd.
Elanco Animal Health A/S
Eli Lilly
Expanscience
Ferring
Fresenius Kabi
Fresenius Medical Care
Galderma
GE Healthcare
Gedeon Richter Nordics AB
Generics Ltd UK
Genzyme Europe B.V.
Gilead
Glenmark Pharmaceuticals
Grünenthal GmbH
GSK
Hospira
Immunocore
Intendis GmbH Berlin
Intercept
Janssen Biologics B.V.
Janssen-Cilag
JSC Grindeks
Kela Laboratories NV
Kyowakirin
LEO Pharma
Les Laboratories Servier
Linde Healthcare – ÍSAGA
Lohmann Animal Health
Lundbeck
LYFIS
Martindale Pharmaceuticals
The Medicines Company UK Limited
MEDA
medac Gesellschaft für klinische Spezialpräparate GmbH
Medical
Medical Valley Invest AB
Medice Arzneimittel Pütter GmbH & Co. KG
Medilink
Medimpex UK Limited
Menarini International Operations Luxembourg S.A.
Mentholatum Company Ltd.
Merck
MercuryPharma
Moderna
MSD
Mirum Pharmaceuticals International B.V.
Mylan AB
Mylan Hospital AS
Mylan SAS
N.V. Organon
Norbrook
Nordic Drugs
Norpharma A/S
Novartis
Novo Nordisk
Octapharma Nordic
Orifarm Generics A/S
Orion Corporation
Otsuka
Pfizer
Pfizer Oy Animal Health
pharma&
PharmaCoDane
Pharmacosmos A/S
Pharma Nord ApS
Pierre Fabré
Raptor Pharmaceuticals Europe B.V.
ratiopharm GmbH
Reckitt Benckiser Healthcare
Recordati AB
Roche
Samsung Bioepis NL B.V
Sandoz GmbH
Sanofi
Sanofi-Pasteur MSD
Santen Oy
Santhera Pharamceuticals
Shire Pharmaceuticals Ltd.
SkyePharma PLC
Smith & Nephew
STADA Arzneimittel AG
Swedish Orphan Biovitrum (Sobi)
Takeda
TEVA
Thea
Therakind Ltd
Tillotts Pharma AB
UCB
United Therapeutics Europe Ltd.
Vétoquinol
Vian S.A.
Vifor Pharma
Vivus BV
Zogenix ROI Ltd.
Williams & Halls ehf.